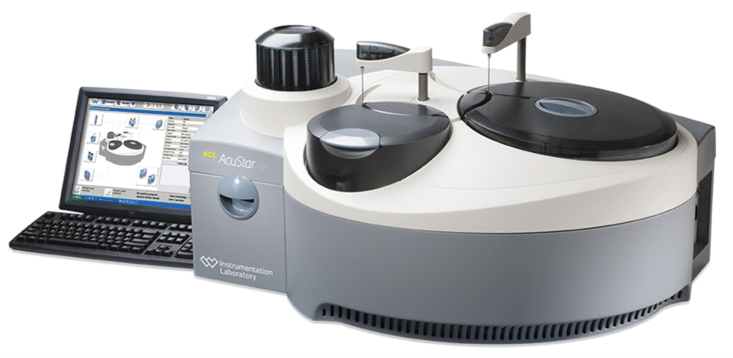
HemosIL AcuStar HIT-IgG(PF4-H). Source: fda.gov
Performance problems could produce false-negative results, potentially delaying diagnosis of a dangerous reaction to heparin.
The U.S. Food and Drug Administration (FDA) has identified a recall of certain lots of the HemosIL AcuStar HIT-IgG(PF4-H) diagnostic test, manufactured by Instrumentation Laboratories (Werfen), as the most serious type of medical device recall.
The recall involves a performance problem that could cause the test to incorrectly report negative results in patients being evaluated for heparin-induced thrombocytopenia (HIT), a potentially life-threatening reaction to the blood-thinning medication heparin.
Why the Recall Matters
Heparin is commonly used in hospitals to prevent or treat blood clots. In some patients, however, it can trigger an immune reaction that reduces platelet counts and increases the risk of dangerous blood clots.
The HemosIL AcuStar HIT-IgG(PF4-H) test helps clinicians evaluate patients for this condition by detecting antibodies associated with HIT.
According to the FDA, customer complaints identified problems with test calibration and inaccurate results. Werfen subsequently confirmed that certain lots could produce results with a negative bias of 20% to 35%, meaning the test could report antibody levels lower than their actual values.
More concerning, the problem may not be reliably detected through routine calibration and quality control testing.
As a result, some patient samples could be incorrectly reported below the test’s cutoff of 1.00 U/mL and interpreted as negative for HIT antibodies.
A false-negative result could delay recognition of HIT and influence treatment decisions. Continued heparin exposure in a patient with HIT could increase the risk of serious thrombotic complications.
Products Affected
The recall affects four lots of HemosIL AcuStar HIT-IgG(PF4-H):
| Part number | Affected lot | Expiration date |
| 0009800127 | B40005 | January 16, 2027 |
| 0009800127 | B38062 | November 1, 2026 |
| 0009802028 | B40006 | January 16, 2027 |
| 0009802028 | B38063 | November 1, 2026 |
Actions for Healthcare Facilities
Werfen has instructed affected customers to identify and destroy remaining products from the recalled lots, document the destruction, and arrange alternative HIT antibody testing methods. The company has also warned that a supply interruption is expected. Werfen’s investigation into the root cause remains ongoing.
Laboratories and treating clinicians should determine whether previously reported patient results require review, follow-up, or repeat testing based on clinical judgment and applicable procedures.
As of September 3, 2026, the company had reported no serious injuries or deaths associated with the issue.
Regulatory Intelligence Perspective
This recall illustrates an important challenge involving diagnostic devices, that a test can produce an apparently valid result even when its analytical performance is compromised. The reported negative bias is particularly concerning because routine calibration and quality control procedures may not reliably identify the problem.
For diagnostic manufacturers and clinical laboratories, the recall underscores the importance of investigating performance complaints, evaluating potential effects on previously reported patient results, and assessing whether existing quality controls are capable of detecting clinically significant failures.
The case also demonstrates why postmarket surveillance and effective corrective action processes are essential throughout a diagnostic device’s commercial life.
Additional information
U.S. customers may contact Werfen Client Services at 1-800-955-9525. FDA recall information is available through the agency’s Medical Device Recalls and Early Alerts website.
FDA Early Alert for Baxter Pressure Breathing Device
FDA Issues Early Alert for Baxter Volara System Patient Circuits Following Therapy Leakage Concerns The U.S. Food and Drug Administration (FDA) has issued an Early Alert regarding a potentially high-risk issue affecting certain Baxter Volara System patient circuits...
FDA Early Alert – Hamilton Medical Breathing Circuit Sets
FDA Issues Early Alert for Hamilton Medical Breathing Circuit Sets Following Reports of Serious Injuries The U.S. Food and Drug Administration (FDA) has issued an Early Alert regarding certain Hamilton Medical coaxial breathing circuit sets used with HAMILTON-C1,...
FDA Early Alert for GE HealthCare Infant Resuscitation Systems
FDA Issues Early Alert for GE HealthCare Infant Resuscitation Systems Due to Oxygen Blender Issue Early Alert for GE HealthCare Infant Resuscitation Systems The U.S. Food and Drug Administration (FDA) has issued an Early Alert regarding a potentially high-risk issue...
FDA-Approved Vivistim: A New Pathway in Stroke Recovery
Vivistim implantable vagus nerve stimulator. Source: Vivistim.com.FDA-Approved Vagus Nerve Stimulation Device Opens a New Pathway in Stroke Recovery A company behind one of the world’s most closely watched neurorehabilitation technologies is preparing for a major new...
FDA Warning Letter Cites Systemic Aseptic Manufacturing Failures
Medline Warning Letter Signals Systemic Quality Failures in Cardiovascular Device Manufacturing The FDA has issued a Warning Letter to Medline Industries (NAMIC Division) following a December 2025 inspection, identifying significant Quality System Regulation (QSR)...
FDA Approves First Gene Therapy for Genetic Hearing Loss
FDA Approves First Gene Therapy for Genetic Hearing LossRapid Review and Breakthrough Science Converge in Landmark OTOF Therapy The FDA has approved Otarmeni (lunsotogene parvec-cwha), the first gene therapy for a form of inherited hearing loss caused by mutations in...
April 2026 Update for FDA QMSR Inspection Results
April 2026 Update for FDA QMSR Inspection ResultsConfirming the Shift to System-Level, Risk-Based Inspection When FDA’s Quality Management System Regulation (QMSR) took effect on February 2, 2026, the regulatory framework became formally aligned with ISO 13485. What...
Kian Pee Wan Capsules Recalled for Undeclared Drug Ingredients
Kian Pee Wan Capsules Recalled Due to Undeclared Drug Ingredients Aphreseller (Buy-herbal.com), an online seller based in Flushing, New York, has voluntarily recalled all lots of Kian Pee Wan capsules after testing identified the presence of undeclared drug...
Good Brain Tonic recalled by Liquid Blenz due to botulism risk
Good Brain Tonic Recall: Liquid Blenz Removes Beverage Due to Botulism Risk FDA Issues Nationwide Alert for “Memory and Focus” Drink Liquid Blenz Corp. has issued a nationwide recall of its Good Brain Tonic beverage due to the potential risk of botulism, a rare but...
Blaine Labs Recalls Wound Care Gels for Microbial Contamination
Blaine Labs, Inc Issues Voluntary Nationwide Recall of Wound Care Gel Products Due to Microbial Contamination Potential infection risk for vulnerable patients highlights critical quality system considerations Recall Overview Blaine Labs, Inc. has issued a voluntary...