Three original device approvals and 218 supplements / notices
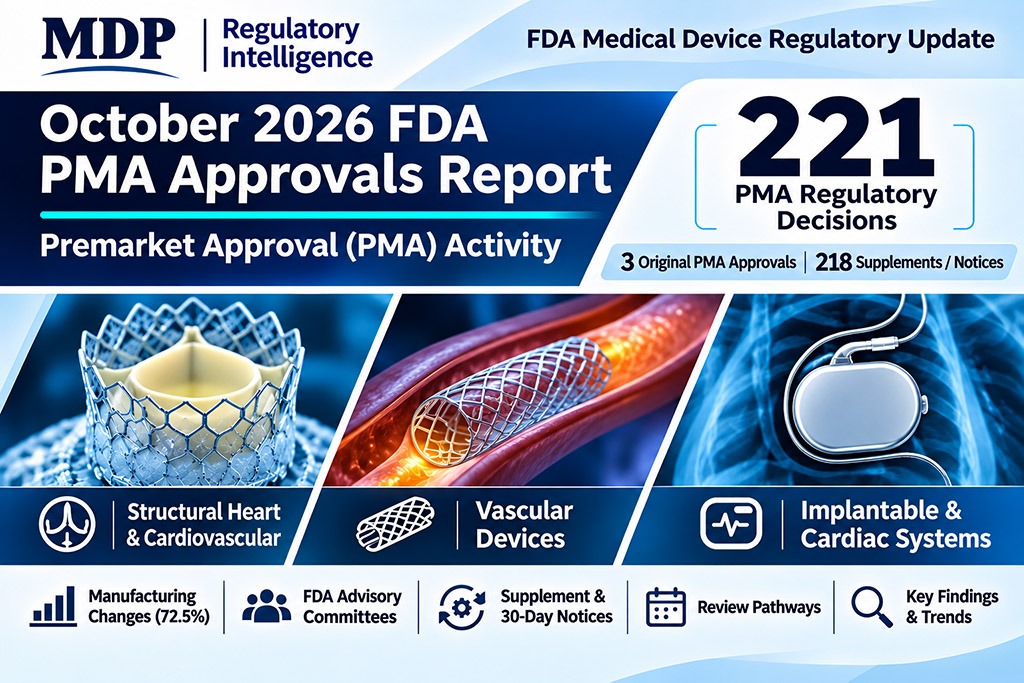
FDA’s October activity highlights three newly approved medical devices and the extensive regulatory oversight required to maintain and modify products already on the market.
The U.S. Food and Drug Administration (FDA) recorded 221 Premarket Approval (PMA) regulatory decisions during October 2026, including three original PMA approvals and 218 supplemental approvals or 30-day notice decisions, according to an analysis by Medical Devices and Pharma (MDP).
Although new medical device approvals often receive the greatest attention, October’s data illustrates another important aspect of medical device regulation: the substantial regulatory activity that continues throughout a device’s commercial life.
MDP’s October 2026 FDA PMA Approvals Report examines original approvals, supplemental submission pathways, FDA advisory committee activity, manufacturing changes, applicant activity, and regulatory decision timing.
Three Medical Devices Receive Original PMA Approval
Three original PMA approvals were recorded during October.
| Manufacturer | Medical device | Approval date |
| Edwards Lifesciences | Autus Size-Adjustable Valve (P260007) | October 1 |
| Vivasure Medical | PerQseal Elite Vascular Closure Device (P250015) | October 16 |
| Abbott Medical | Amplatzer Paraclose PVL Occluder (P260019) | October 27 |
These approvals represent new authorizations through FDA’s PMA pathway, generally applicable to Class III medical devices requiring the highest level of regulatory review.
Unlike the 510(k) pathway, which generally relies on demonstrating substantial equivalence to a legally marketed predicate device, PMA requires valid scientific evidence providing reasonable assurance of safety and effectiveness for the device’s intended use.
The three October approvals illustrate continued innovation in cardiovascular and related medical technologies.
Most PMA Activity Involves Previously Approved Devices
Of the 221 October decisions, 218 involved PMA supplements or 30-day notices, representing approximately 98.6% of all recorded activity.
The distribution of regulatory pathways included:
| PMA pathway | Decisions |
| 30-Day Notice | 150 |
| Real-Time Process | 22 |
| Normal 180-Day Track — No User Fee | 19 |
| Normal 180-Day Track | 13 |
| Special (Immediate Track) | 6 |
| 135 Review Track | 5 |
| Panel Track | 3 |
| Total supplements and notices | 218 |
The three original PMA approvals bring the monthly total to 221. Notably, 30-day notices accounted for 67.9% of all October PMA decisions.
These notices are generally associated with certain manufacturing changes that affect the safety or effectiveness of an approved device. They are distinct from original PMA approvals and should not be interpreted as newly authorized medical devices.
Manufacturing and Process Changes Dominate Regulatory Activity
Perhaps the most significant finding concerns the reasons manufacturers submitted PMA supplements and notices. Of the 218 supplemental and notice decisions, 158 involved manufacturing, sterilization, packaging, or supplier process changes. That represents 72.5% of all supplement and notice activity.
Other regulatory changes included:
- Design, component, specification, or material changes: 28 decisions.
- Postapproval study-related activity: 14 decisions.
- Labeling changes: 13 decisions.
- Manufacturing location changes: four decisions.
- Postapproval study-related labeling: one decision.
These findings highlight the extensive regulatory responsibilities associated with maintaining approved medical devices. Manufacturers must continually evaluate changes to production processes, materials, suppliers, equipment, and manufacturing facilities to determine whether FDA notification or approval is required.
For quality and regulatory professionals, this activity underscores the importance of effective design and process change control, supplier management, risk management, and regulatory impact assessments. A device’s original PMA approval establishes its authorization for marketing, but subsequent changes can create additional regulatory obligations throughout its lifecycle.
Cardiovascular Devices Account for More Than Half of PMA Decisions
October’s PMA activity was concentrated in several medical specialties.
| FDA advisory committee | Decisions | Share |
| Cardiovascular | 113 | 51.1% |
| Neurology | 30 | 13.6% |
| Ophthalmic | 18 | 8.1% |
| Gastroenterology/Urology | 14 | 6.3% |
| General & Plastic Surgery | 11 | 5.0% |
| Other committees | 35 | 15.8% |
Cardiovascular devices accounted for more than half of all PMA decisions, reflecting substantial regulatory activity involving cardiac implants, vascular devices, and related technologies. Neurology and ophthalmic devices were also prominent.
These figures reflect FDA advisory committee classifications and include both original approvals and lifecycle decisions. They should not be interpreted as the number of newly approved devices within each specialty.
Regulatory Activity Extends Across Major Manufacturers
October’s dataset included activity involving established medical device manufacturers such as Biotronik, Abbott, Medtronic, and Edwards Lifesciences.
Many decisions involved modifications to existing device families rather than entirely new technologies. This distinction is important when interpreting manufacturer activity. A company with numerous PMA supplement decisions is not necessarily introducing an equivalent number of new devices.
Instead, the activity may reflect ongoing improvements, manufacturing modifications, supplier changes, or other regulatory maintenance of existing product portfolios.
Understanding PMA Review Timing
MDP also examined elapsed calendar time between FDA’s recorded receipt and decision dates for the different PMA submission pathways. These measurements provide insight into the timing associated with regulatory decisions, but they should not be confused with official FDA review-clock performance.
Elapsed calendar time may include periods when FDA is awaiting additional information from manufacturers, administrative processing, or other activities not counted toward statutory review goals.
The report provides additional analysis of decision timing across the various PMA pathways.
What October’s PMA Activity Tells Us
October’s results demonstrate that FDA’s PMA program involves considerably more than evaluating new Class III medical devices.
While three original approvals introduced newly authorized technologies, the overwhelming majority of recorded regulatory decisions involved previously approved products.
The prominence of manufacturing and process changes also illustrates the close relationship between regulatory affairs, quality management, manufacturing operations, and supplier oversight.
For medical device manufacturers, this reinforces a fundamental principle. FDA approval is not the end of the regulatory process. It is the beginning of an ongoing responsibility to maintain device safety, effectiveness, manufacturing controls, and regulatory compliance.
Understanding these lifecycle obligations is essential to sustaining market authorization and managing changes to approved medical devices.
MDP’s comprehensive report provides additional information and analysis, including original PMA approvals and device details, PMA supplement and 30-day notice pathways, FDA advisory committee distribution, manufacturing and regulatory change categories, manufacturer activity, regulatory decision timing, and noteworthy findings and regulatory intelligence observations.
Source: FDA Premarket Approval database, October 2026 decision records. MDP analysis reflects decisions recorded during the reporting period and may be subject to subsequent FDA database updates. This is an independent MDP Regulatory Intelligence publication and is not an official FDA report.
FDA Approves Novel Device for Pancreatic Cancer Treatment
FDA Approves Novel Device for Pancreatic Cancer Treatment The U.S. Food and Drug Administration has approved Optune Pax, a first-of-its-kind medical device designed to treat adult patients with locally advanced pancreatic cancer. Developed by Novocure, the portable...
FDA “PreCheck” Program to Boost U.S. Drug Manufacturing
FDA Announces New PreCheck Drug Manufacturing Program The U.S. Food and Drug Administration (FDA) has announced the launch of FDA PreCheck, a groundbreaking new program designed to strengthen America’s domestic pharmaceutical supply chain by streamlining the...
What Are Early Recall Alerts?
Recall Communication “Early Alerts” Note: This summary is based on an FDA communication attributed to Michelle Tarver, M.D., Ph.D., director of the Center for Devices and Radiological Health (CDRH). In November 2024 the U.S. Food and Drug Administration’s Center for...
The FDA Orphan Drug Program
Incentives for Rare Disease Drug Development The FDA’s Orphan Drug Program was established to encourage the development of drugs for rare diseases that affect fewer than 200,000 people in the U.S. Because these diseases often lack market incentives due to small...
FDA Grants Accelerated Approval to Multiple Myeloma Treatment
For hard-to-treat multiple myeloma On July 2, 2025, the FDA granted accelerated approval to linvoseltamab-gcpt (brand name: Lynozyfic) for adults with relapsed or refractory multiple myeloma who have received at least four prior therapies, including: A proteasome...
FDA Approves Alhemo for Patients with Hemophilia A or B
For Patients with Hemophilia A or B with Inhibitors The FDA has approved Alhemo (concizumab-mtci) for routine prophylaxis to prevent or reduce bleeding episodes in patients aged 12 and older with: Hemophilia A with factor VIII inhibitors, or Hemophilia B with factor...
First-in-Class Non-Opioid Pain Reliever: Journavx
A new non-opioid pain treatment option On January 30, 2025, the FDA granted priority review for Journavx (suzetrigine) – a first-in-class, non-opioid analgesic for treating moderate to severe acute pain in adults. How It Works Journavx targets sodium channels in the...
FDA Approves DATROWAY® Breast Cancer Treatment
A new breast cancer treatment option The United States Food and Drug Administration (FDA) approved DATROWAY® (datopotamab deruxtecan-dlnk) on January 17, 2025, for adults with hormone receptor-positive (HR+), HER2-negative breast cancer that is unresectable or...
Human Factors Engineering
Human Factors Engineering Human Factors Engineering (HFE) is critical for medical device safety and regulatory compliance, especially for Class II and III devices, drug-delivery systems, and combination products. Following is a detailed breakdown of the regulatory...
Generic Eliquis
Generic Eliquis Editor's note: FDA originally announced the approval of generic equivalents to Eliquis in 2019, and updated that information the next year. We are publishing this information as a response to reader requests for more information about generic Eliquis....