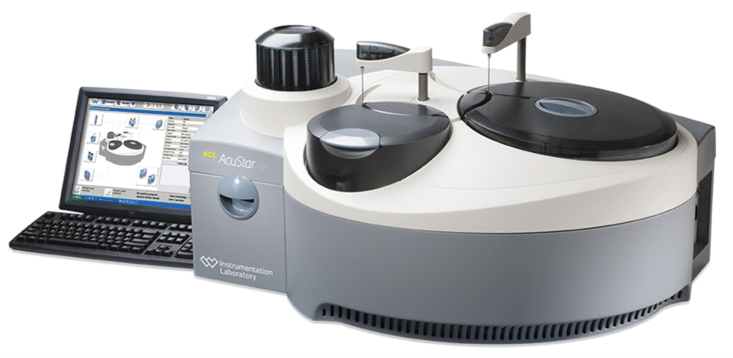
HemosIL AcuStar HIT-IgG(PF4-H). Source: fda.gov
Performance problems could produce false-negative results, potentially delaying diagnosis of a dangerous reaction to heparin.
The U.S. Food and Drug Administration (FDA) has identified a recall of certain lots of the HemosIL AcuStar HIT-IgG(PF4-H) diagnostic test, manufactured by Instrumentation Laboratories (Werfen), as the most serious type of medical device recall.
The recall involves a performance problem that could cause the test to incorrectly report negative results in patients being evaluated for heparin-induced thrombocytopenia (HIT), a potentially life-threatening reaction to the blood-thinning medication heparin.
Why the Recall Matters
Heparin is commonly used in hospitals to prevent or treat blood clots. In some patients, however, it can trigger an immune reaction that reduces platelet counts and increases the risk of dangerous blood clots.
The HemosIL AcuStar HIT-IgG(PF4-H) test helps clinicians evaluate patients for this condition by detecting antibodies associated with HIT.
According to the FDA, customer complaints identified problems with test calibration and inaccurate results. Werfen subsequently confirmed that certain lots could produce results with a negative bias of 20% to 35%, meaning the test could report antibody levels lower than their actual values.
More concerning, the problem may not be reliably detected through routine calibration and quality control testing.
As a result, some patient samples could be incorrectly reported below the test’s cutoff of 1.00 U/mL and interpreted as negative for HIT antibodies.
A false-negative result could delay recognition of HIT and influence treatment decisions. Continued heparin exposure in a patient with HIT could increase the risk of serious thrombotic complications.
Products Affected
The recall affects four lots of HemosIL AcuStar HIT-IgG(PF4-H):
| Part number | Affected lot | Expiration date |
| 0009800127 | B40005 | January 16, 2027 |
| 0009800127 | B38062 | November 1, 2026 |
| 0009802028 | B40006 | January 16, 2027 |
| 0009802028 | B38063 | November 1, 2026 |
Actions for Healthcare Facilities
Werfen has instructed affected customers to identify and destroy remaining products from the recalled lots, document the destruction, and arrange alternative HIT antibody testing methods. The company has also warned that a supply interruption is expected. Werfen’s investigation into the root cause remains ongoing.
Laboratories and treating clinicians should determine whether previously reported patient results require review, follow-up, or repeat testing based on clinical judgment and applicable procedures.
As of September 3, 2026, the company had reported no serious injuries or deaths associated with the issue.
Regulatory Intelligence Perspective
This recall illustrates an important challenge involving diagnostic devices, that a test can produce an apparently valid result even when its analytical performance is compromised. The reported negative bias is particularly concerning because routine calibration and quality control procedures may not reliably identify the problem.
For diagnostic manufacturers and clinical laboratories, the recall underscores the importance of investigating performance complaints, evaluating potential effects on previously reported patient results, and assessing whether existing quality controls are capable of detecting clinically significant failures.
The case also demonstrates why postmarket surveillance and effective corrective action processes are essential throughout a diagnostic device’s commercial life.
Additional information
U.S. customers may contact Werfen Client Services at 1-800-955-9525. FDA recall information is available through the agency’s Medical Device Recalls and Early Alerts website.
Early Alert: Cook Medical Sizing Catheters
Cook Medical Sizing Catheters Removed Due to Risk of Device Breakage The U.S. Food and Drug Administration (FDA) has issued an early alert regarding a potentially serious device issue involving certain sizing catheters manufactured by Cook Medical. Cook Medical has...
Medline Recalls Angiographic RA Control Syringes
Angiographic Syringe Recall: Medline Removes RA Control Syringes Due to Risk of Disconnection and Air Embolism The U.S. Food and Drug Administration (FDA) has announced that Medline Industries has issued an Urgent Medical Device Recall involving certain Namic...
FDA Class I Recall for Abiomed Impella Gen 1 Purge Cassettes
Medical Device Alert: Abiomed Impella Purge Cassette Recall (Class I) Abiomed has initiated a Class I recall, the most serious classification issued by the U.S. Food and Drug Administration, for certain Impella purge cassettes and associated pump sets due to a...
When a Small Process Failure Becomes a Patient Safety Event
What the Erbe Cryoprobe Alert Teaches Us About Modern Device Quality The FDA’s recent Early Alert regarding flexible cryoprobes manufactured by Erbe USA is a textbook example of a fundamental truth in medical device quality. That small manufacturing deviations can...
EPA Proposes Rollback of EtO Sterilization Emissions Limits
EPA Proposes Rollback of Ethylene Oxide Sterilization Emissions LimitsThe U.S. Environmental Protection Agency has proposed weakening restrictions on emissions of ethylene oxide (EtO) from commercial sterilization facilities that sterilize medical devices. EtO plays a...
Adulterated Supplements and the Limits of Post-Market Oversight
FDA Recall of Adulterated Supplements Raises Questions About DSHEA Oversight Lessons from the “Boner Bear” Recall The FDA’s recent recall of several honey-based “sexual enhancement” supplements illustrates a recurring problem in the dietary supplement marketplace:...
FDA Recalls “Sexual Enhancement” Supplements
FDA Recall Highlights Ongoing Risks of Adulterated “Sexual Enhancement” Supplements Pure Vitamins and Natural Supplements, LLC (Tampa, FL) has issued a voluntary nationwide recall of several honey-based dietary supplement products after FDA laboratory testing detected...
When Apparel Becomes a Medical Device
Lessons from FDA’s Breast Binder Warning Letters In December 2025, the FDA issued warning letters to several companies selling breast binders and chest compression garments online. The agency concluded that marketing claims on company websites, like statements about...
Ethicon Issues Safety Alert for Surgical Stapler Cartridge
Endopath Echelon Vascular White Reload for Advanced Placement TipSource: FDAUrgent Class I Medical Device Recall Ethicon Endo-Surgery, LLC has issued an urgent correction for its Endopath Echelon Vascular White Reload for Advanced Placement Tip (35 mm, 4-row). This...
FDA Cracks Down on Insightra Medical
Warning Letter Cites a Number of Serious Violations In a scathing warning letter, the FDA has cited Insightra Medical, Inc. for widespread regulatory violations involving its Intra-Aortic Balloon Pump (IABP) Catheter Kit—a device used in critical cardiovascular...